GPCR-AI DISCOVERY PLATFORM
From receptor dynamics to therapeutic candidates.
Starting from molecular mechanism, we connect structure, computation, artificial intelligence and functional experiments in a rapid iterative system.
02 / 06
PLATFORM LOGIC
Prediction and experiment calibrate each other in every design cycle.
We do not treat AI as an isolated tool. It is embedded across GPCR mechanism research, structural analysis, molecular generation, virtual screening, property optimization and biological testing.
UNDERSTANDDESIGNTEST
 01
SCIENCE AREA 01
GPCR molecular research
Membrane proteins
in particular G protein-coupled receptors (GPCRs), are the most important drug targets. As an example, about 40% of the clinically used drugs are targeting GPCRs.
In some pharmaceutical companies, this figure can be even as high as 70-80%. Therefore, the understanding of membrane protein functions is of fundamental importance for modern drug discovery.
In some pharmaceutical companies, this figure can be even as high as 70-80%. Therefore, the understanding of membrane protein functions is of fundamental importance for modern drug discovery.
AlphaMol
Our team members have made important contributions to understand receptor mediated signalling at the molecular level. They have published many innovative works in top journals including Cell, Nature,
Nature Communications, Chemical Science, Trends in Biochemical Sciences, Angewandte Chemie and others.
Nature Communications, Chemical Science, Trends in Biochemical Sciences, Angewandte Chemie and others.
Publication18 selected works
Functional dynamics of G protein-coupled receptors reveal new routes for drug discoveryNature Reviews Drug Discovery
The structure and function of olfactory receptorsTrends in Pharm Sci
Molecular basis of ligand selectivity for melatonin receptorsRSC Adv
The role of metal ions in GPCR signaling and drug discoveryComputational Molecular Science
Structural basis of CXC chemokine receptor 2 activation and signallingNature
Activation and Signaling Mechanism Revealed by Cannabinoid Receptor-Gi Complex StructuresCell
Enhancing the Signaling of GPCRs via Orthosteric IonsACS Central Science
New binding sites, new opportunities for GPCR drug discoveryTrends in Biochemical Sciences
New binding sites, new opportunities for GPCR drug discoveryTrends in Biochemical Sciences
Exploring a new ligand binding site of G protein-coupled receptorsChemical Science
5-HT2C Receptor Structures Reveal the Structural Basis of GPCR PolypharmacologyCell
Designing Safer Analgesics via µ-Opioid Receptor PathwaysTrends in Pharmacological Sciences
The Molecular Mechanism of P2Y1 Receptor ActivationAngewandte Chemie Int Ed Engl
Mechanistic Studies on the Stereoselectivity of the Serotonin 5-HT1A ReceptorAngewandte Chemie Int Ed Engl
The Mechanism of Ligand-Induced Activation or Inhibition of mu- and kappa-Opioid ReceptorsAngewandte Chemie Int Ed Engl
W246 Opens a Gate for a Continuous Intrinsic Water Pathway during Activation of the Adenosine A ReceptorAngewandte Chemie Int Ed Engl
Activation of G-protein-coupled receptors correlates with the formation of a continuous internal water pathwayNature Communications
Advances in GPCR modeling evaluated by the GPCR Dock 2013 assessment: meeting new challengesStructure
 02
SCIENCE AREA 02
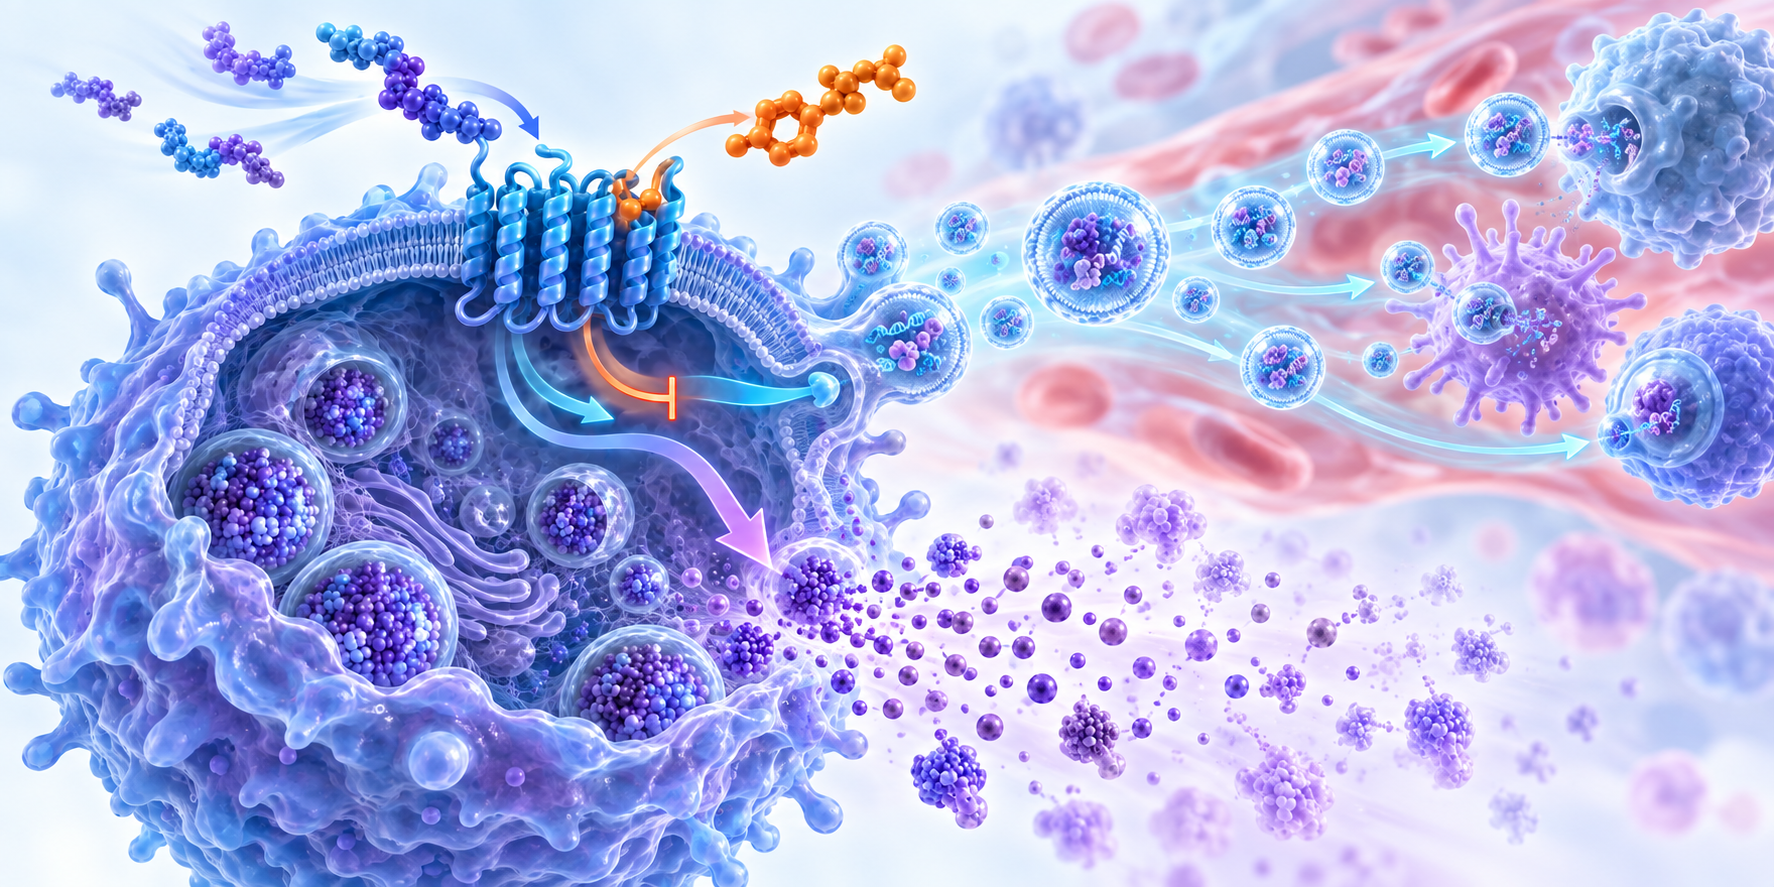
Innovative GPCR biotesting
G protein-coupled receptors
G protein-coupled receptors (GPCRs) are the most important drug targets, of which about 40% of the marketed drugs are targeting GPCRs.
As a membrane protein, the biological activity testing for GPCR drug discovery is still a challenging task. Many difficulties remain to be resolved including: (1) testing drug activity without labeling (2) measuring GPCR/G-protein interaction directly instead of determining that of cAMP or Ca2+ signaling in an indirect way (3) measuring kinetic properties of GPCR drug molecules and (4) differentiating G protein types inside cells.
As a membrane protein, the biological activity testing for GPCR drug discovery is still a challenging task. Many difficulties remain to be resolved including: (1) testing drug activity without labeling (2) measuring GPCR/G-protein interaction directly instead of determining that of cAMP or Ca2+ signaling in an indirect way (3) measuring kinetic properties of GPCR drug molecules and (4) differentiating G protein types inside cells.
In this regard, the team member of AlphaMol developed a series of innovative biochip technologies to overcome the hurdle in GPCR drug discovery in the past 20 years. Such breakthrough facilitates the process of GPCR drug discovery in a much more accurate and efficient way.
Publication4 selected works
Single-vesicle assays using liposomes and cell-derived vesicles: from modeling complex membrane processes to synthetic biology and biomedical applicationsChemical reviews
Activation of G-protein-coupled receptors in cell-derived plasma membranes supported on porous beadsJACS
Micropatterned immobilization of a G protein–coupled receptor and direct detection of G protein activationNature Biotechnology
Incorporation of Rhodopsin in Laterally Structured Supported Membranes: Observation of Transducin Activation with Spatially and Time-Resolved Surface Plasmon Resonance†Biochemistry
 03
SCIENCE AREA 03
New drug discovery
AlphaMol
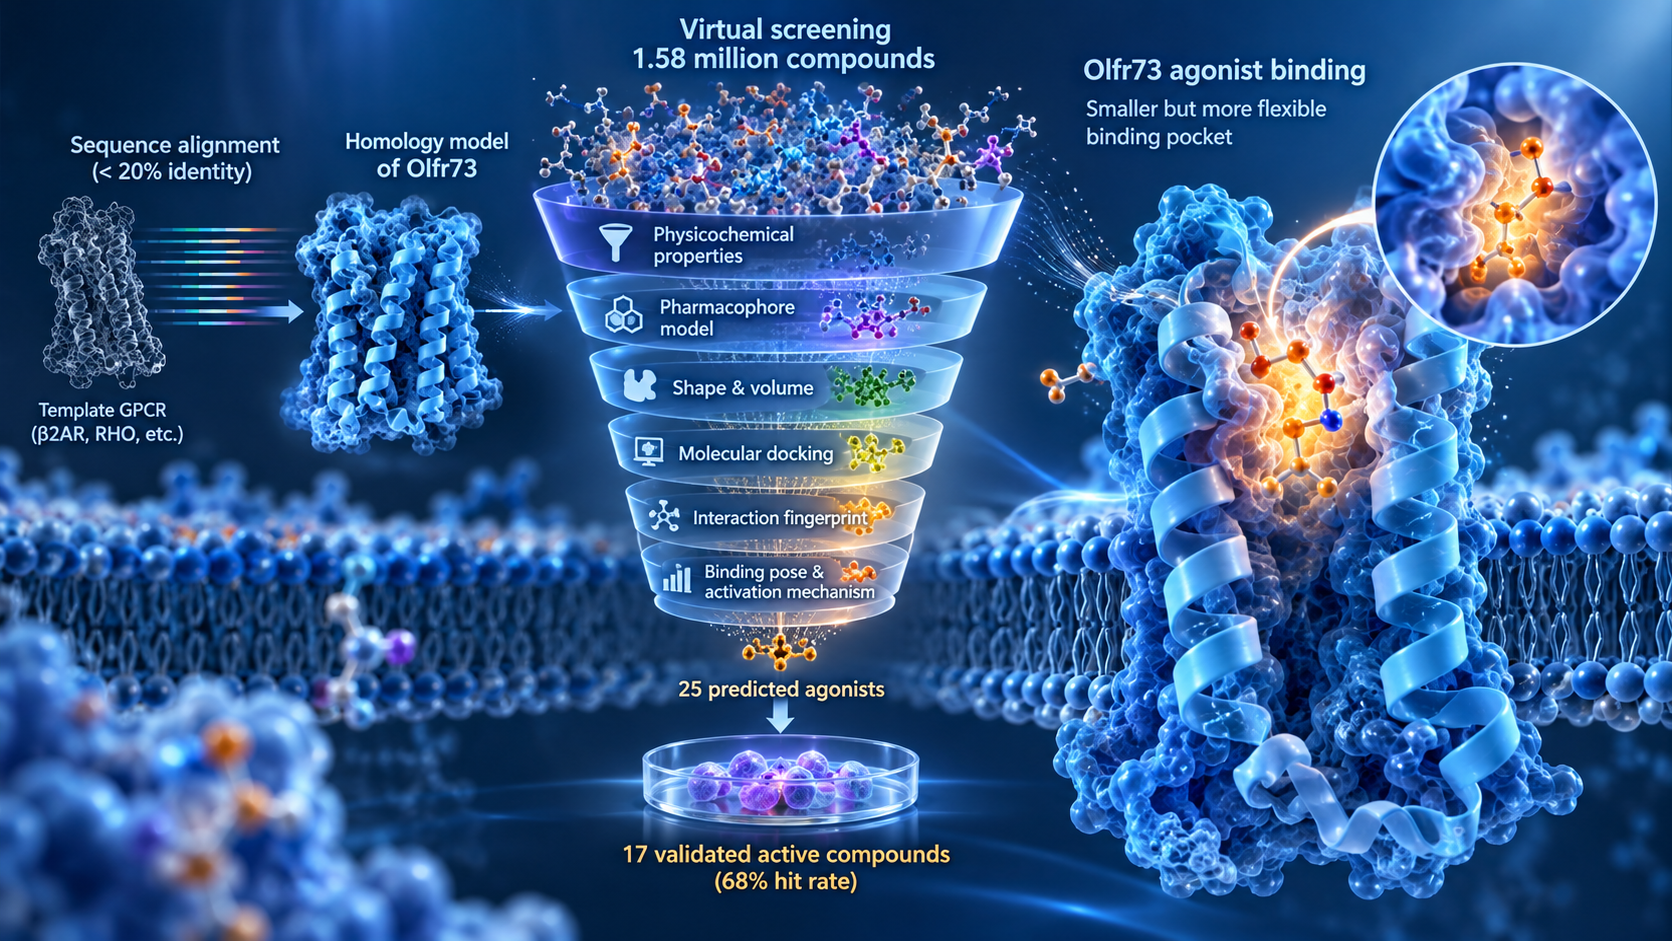
Modern drug discovery and development is a complicated process, which costs about $2-3 billion and lasts for 12 years on average. How to decrease the costs and speed up the process is the challenge. Computational biology and Artificial Intelligence, combined with new technologies in experimental drug screening, are expected to make the hunt for new drugs quicker, cheaper and more efficient.
Our team has established a sophisticated computer-aided drug discovery platform, which combines Artificial Intelligence, WebGL, cloud computing and big data. It can perform a wide range of drug discovery tasks such as molecular generation, virtual screening, lead optimization, drug property optimization, molecular dynamics simulation, accurate binding energy prediction and others. We have applied our tools successfully to discover and design potent drug candidates for various targets.
Members of our team have the experience of advancing designed molecules into clinical trials. More recently, we discovered a series of potent compounds for different orphan receptors. These compounds could result in "me-only" drug candidates.
Our team has established a sophisticated computer-aided drug discovery platform, which combines Artificial Intelligence, WebGL, cloud computing and big data. It can perform a wide range of drug discovery tasks such as molecular generation, virtual screening, lead optimization, drug property optimization, molecular dynamics simulation, accurate binding energy prediction and others. We have applied our tools successfully to discover and design potent drug candidates for various targets.
Members of our team have the experience of advancing designed molecules into clinical trials. More recently, we discovered a series of potent compounds for different orphan receptors. These compounds could result in "me-only" drug candidates.
Publication17 selected works
Enhancing Multifunctional Drug Screening via Artificial IntelligenceDigital Discovery
Identifying Potent Compounds Using Pairwise Consensus MethodsJournal of Chemical Information and Modeling
Rational Design Biased Compounds against μ-Opioid ReceptorACS Pharmacology & Translational Science
Functional dynamics of G protein-coupled receptors reveal new routes for drug discoveryNature Reviews Drug Discovery
Computational drug development for membrane protein targetsNature Biotechnology
Will the hype of automated drug discovery finally be realized?Expert Opinion on Drug Discovery
Celastrol Combats Methicillin-Resistant Staphylococcus aureus by Targeting Δ1-Pyrroline-5-Carboxylate DehydrogenaseAdvanced Science
Rutin, a Natural Inhibitor of IGPD Protein, Inhibits the Formation of Biofilm in Staphylococcus xylosus ATCC700404 in vitro and in vivoFrontiers in Pharmacology
MolADI: A Web Server for Automatic Analysis of Protein–Small Molecule Dynamic InteractionsMolecules
Clinical HDAC Inhibitors are Effective Drugs to Prevent the Entry of SARS-CoV2ACS Pharmacology & Translational Science
Enhancing the Signaling of GPCRs via Orthosteric IonsACS Central Science
Advancing Drug Discovery via Artificial IntelligenceTrends in Pharmacological Sciences
Computational modeling of the olfactory receptor Olfr73 suggests a molecular basis for low potency of olfactory receptor-activating compoundsCommunications Biology
New binding sites, new opportunities for GPCR drug discoveryTrends in Biochemical Sciences
Exploring a new ligand binding site of G protein-coupled receptorsChemical Science
Implementing WebGL and HTML5 in Macromolecular Visualization and Modern Computer-Aided Drug DesignTrends in Biotechnology
Using PyMOL as a platform for computational drug designComputational Molecular Science
 04
SCIENCE AREA 04
Resolving membrane protein structures in real lipids
Membrane proteins
Membrane proteins are important drug targets which include G protein-coupled receptors (GPCRs), ion channels, transporters and others. Our team has close collaborations with leading scientist in the area of structural biology. However, most of current membrane protein structures were resolved at detergent or artificial lipid environments which are totally different from their physiological conditions. This in return result in losing the physiological functions of target membrane protein. Such structures will increase the cost the of structure-based drug discovery noticeably.
AlphaMol
Our team has developed a new pipeline which can resolve the membrane protein structures at real lipid environments. With such functional structures, we can design drug molecules in a more accurate way. Moreover, excellent cryoEM facilities are accessible in both China and Switzerland.
Publication8 selected works
Cryo-EM structures of ClC-2 chloride channel reveal the blocking mechanism of its specific inhibitor AK-42Nat Commun
Cryo-EM structure of human heptameric pannexin 2 channelNature Communications
Asymmetric opening of the homopentameric 5-HT3A serotonin receptor in lipid bilayersNature Communications
A Gating Mechanism of the Serotonin 5-HT3 ReceptorStructure
The Structure of the Mouse Serotonin 5-HT3 Receptor in Lipid VesiclesStructure
X-ray structure of the mouse serotonin 5-HT3 receptorNature
Molecular and dimensional profiling of highly purified extracellular vesicles by fluorescence fluctuation spectroscopyAnalytical Chemistry
Overcoming barriers to membrane protein structure determinationNature Biotechnology
 05
SCIENCE AREA 05
New computational algorithm development
AlphaMol
Our team has developed an absolute binding energy prediction tool, which is known as AlphaE. With the new algorithm of AlphaE, we can predict accurately the binding activity of a compound. The RMSE (root-mean-square-energy) is as low as 1 kcal/mol in comparison with experimental data. AlphaE also overcomes many difficulties in currently available tools such as: (1) predictions are not reliable when the molecule changes too much; (2) calculations cannot be performed if molecules bind to different pockets; (3) predictions need large amount of CPU and GPU resources.
Furthermore, AlphaMol also developed many new computational algorithm and tools to predict the physical properties and drug-likeness of molecules. These tools demonstrated perfect correlations with experimental data, of which the R2 is up to 0.99.
Furthermore, AlphaMol also developed many new computational algorithm and tools to predict the physical properties and drug-likeness of molecules. These tools demonstrated perfect correlations with experimental data, of which the R2 is up to 0.99.
Publication6 selected works
Machine learning deciphered molecular mechanistics with accurate kinetic and thermodynamic predictionJournal of Chemical Theory and Computation
Accurate Physical Property Predictions via Deep LearningMolecules
MolADI: A Web Server for Automatic Analysis of Protein–Small Molecule Dynamic InteractionsMolecules
Advancing Drug Discovery via Artificial IntelligenceTrends in Pharmacological Sciences
Implementing WebGL and HTML5 in macromolecular visualization and modern computer-aided drug designTrends in Biotechnology
Using PyMOL as a platform for computational drug designComputational Molecular Science
DISCOVER TOGETHER
Working on a difficult GPCR problem?
Explore a collaboration with our scientific team.